
KRAS G12D Mutation: From an "Undruggable" Target to Breakthroughs in PROTAC Degradation Strategies
Author: Senior Product Manager of UA Views:11 


This article focuses on the molecular characteristics and targeted therapeutic strategies of the KRAS G12D mutation, systematically elucidating the distribution patterns of RAS family gene mutations and their molecular mechanisms driving tumorigenesis. It provides an in-depth analysis of the challenges in drug development for KRAS G12D due to the lack of covalent binding sites and explores the novel approach of VHL-based PROTAC technology for targeted degradation of KRAS G12D.
- Recent Advances
- Product Information
Recent Advances
KRASG12D Mutation: From an "Undruggable" Target to the Breakthrough of PROTAC Degradation Strategy
Summary
This article systematically explores the molecular characteristics of KRASG12D mutation and its targeted therapeutic strategies, detailing the distribution patterns of RAS family gene mutations and their mechanisms in driving tumorigenesis. It analyzes the challenges in drug development for KRASG12D due to the lack of covalent binding sites and discusses the new pathways provided by VHL-based PROTAC technology for targeted degradation of KRASG12D.
This article systematically explores the molecular characteristics of KRASG12D mutation and its targeted therapeutic strategies, detailing the distribution patterns of RAS family gene mutations and their mechanisms in driving tumorigenesis. It analyzes the challenges in drug development for KRASG12D due to the lack of covalent binding sites and discusses the new pathways provided by VHL-based PROTAC technology for targeted degradation of KRASG12D.
1. RAS Gene Family and Clinical Distribution of KRAS Mutations
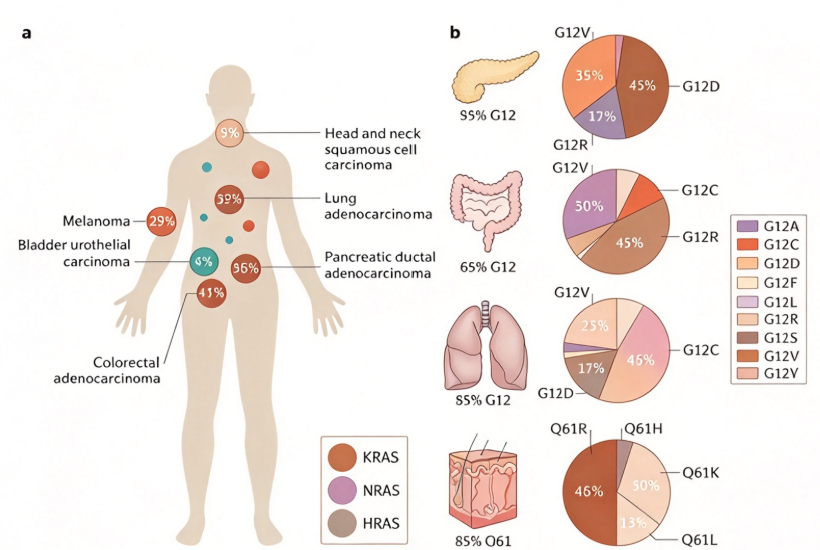
RAS, the first identified human proto-oncogene, is the most widely mutated oncogene in tumors, appearing in nearly 30% of human cancers. The RAS family includes three subtypes: KRAS, HRAS, and NRAS, encoding four proteins: HRAS, NRAS, KRAS4A, and KRAS4B. KRAS mutations account for approximately 85% of RAS family mutations and are more likely to drive tumorigenesis compared to other RAS family gene mutations.
KRAS functions as a molecular switch, transitioning between the GTP-bound active state and the GDP-bound inactive state to transmit signals from membrane-bound receptors and regulate cellular processes. KRAS mutations primarily occur at four hotspot codons (12, 13, 61, and 146). Codon 12 is the most frequently mutated among these hotspots, with G12D being the most common, followed by G12V, G12C, and others. KRAS mutations and/or wild-type KRAS amplification are prevalent in pancreatic cancer, colorectal cancer, and non-small cell lung cancer. In the U.S., G12C is mutated in 13.6% of lung adenocarcinomas, while G12D and G12V are the two most common mutations in colorectal and pancreatic cancers.
2. Molecular Basis of KRAS's "Undruggable" Nature
KRAS has been dubbed the "Death Star" protein. Since its discovery in 1982, it has long been considered "undruggable." The primary reasons include: First, RAS has picomolar affinity for GTP, while intracellular GTP concentrations reach 0.5 mmol/L, making the development of competitive inhibitors extremely challenging. Second, the KRAS protein surface is smooth, with a nearly spherical spatial structure lacking small-molecule binding pockets. Third, the D-domains of RAS subtypes are highly similar, resulting in poor targeting selectivity.
The discovery of the switch pocket II in KRAS protein and the newly introduced cysteine residue at codon 12 in KRASG12C mutation, which readily forms covalent bonds, led to the design of covalent small-molecule inhibitors that irreversibly target and bind to the cysteine residue at codon 12. The successful market approval of KRAS G12C-targeted inhibitors demonstrated that KRAS mutations are not "undruggable." This breakthrough has injected confidence into the field of KRAS-targeted drug development and provided important references for exploring strategies targeting other mutant subtypes.
3. Unique Challenges in Drug Development for KRASG12D Mutation
KRAS G12C mutation has been clinically proven to be directly inhibited by small-molecule covalent inhibitors, which trap the protein in the inactive GDP-bound state. However, the GTP hydrolysis rate of KRAS G12D is 2 to 3 times lower than that of KRAS G12C, leading to more constitutive activation. Therefore, despite the presence of similar binding pockets in the GDP state, pharmacology targeting the inactive state is unlikely to achieve results comparable to those for G12C.
Additionally, the KRAS G12D mutation involves the substitution of glycine at codon 12 with aspartate, which terminates in a carboxyl group. This mutant subtype lacks amino acid residues near switch pocket II for covalent binding, making selective covalent binding strategies unfeasible. Thus, new methods are required to develop selective inhibitors with high affinity and drug-like potency. Targeted drug development for KRAS G12D mutation has become one of the most closely watched directions. KRAS G12D is the most prevalent KRAS mutation and the most common KRAS mutant subtype in pancreatic and colon cancers. Pancreatic cancer, often referred to as the "king of cancers," has limited treatment options, making drug development for this target an urgent clinical need.
4. PROTAC Technology Offers a New Pathway for Targeted Degradation of KRASG12D
Proteolysis-targeting chimeras (PROTAC) technology leverages the ubiquitin-proteasome system within cells to deliver target proteins to the proteasome for degradation. Unlike traditional small-molecule inhibitors that block protein function by occupying active sites, PROTAC molecules bind the target protein at one end and recruit E3 ligases at the other, forming a ternary complex of target protein-PROTAC-E3 ligase. This mediates the polyubiquitination of the target protein and its subsequent degradation by the proteasome.
For the KRASG12D mutation, PROTAC technology provides a strategy that does not rely on covalent binding. By designing target protein ligands that bind KRASG12D with high affinity and recruiting E3 ligases such as VHL, selective degradation of KRASG12D protein can be achieved. VHL-based PROTAC molecules have been successfully applied in degradation studies of various target proteins, and their application in KRASG12D degradation represents a significant direction in this field. The ability to form ternary complexes is a critical parameter determining the degradation efficiency of PROTACs, making it essential to establish efficient and reliable binding detection methods at this stage.
5. Conclusion
In KRAS G12D-targeted therapy research, in addition to PROTAC degradation strategies, T-cell immune-based targeting strategies have also garnered widespread attention. The abnormal protein produced by the KRAS G12D mutation, after degradation by the proteasome, generates short peptide fragments containing the mutation site (e.g., VVVGADGVGK). These fragments are presented on the tumor cell surface by HLA molecules, forming neoantigens recognizable by T cells. To meet the demand for detecting KRAS G12D-specific T cells, UA offers the UA-MHC HLA-A1101/VVVGADGVGK KRAS G12D Tetramer-APC Labelled. This product is an APC-labeled tetramer complex that specifically identifies HLA-A1101-restricted KRAS G12D neoantigen-specific CD8+ T cells. It is suitable for screening and validating KRAS G12D-specific TCRs, quality control of TCR-T cell products, and detecting antigen-specific T cells in tumor immunotherapy research.
As the most prevalent KRAS mutant subtype, KRAS G12D faces drug development challenges due to the lack of covalent binding sites, but these are gradually being overcome by emerging technologies. VHL-based PROTAC technology, by inducing selective degradation of KRAS G12D protein, provides a novel intervention strategy for this "undruggable" target. Meanwhile, T-cell immunotherapy strategies based on KRAS G12D neoantigens are also advancing. The UA-MHC HLA-A*1101/VVVGADGVGK KRAS G12D Tetramer-APC Labelled offers a reliable tool for detecting KRAS G12D-specific T cells and developing TCR-T cell therapies, continuously driving innovation in the field of KRAS G12D-targeted therapy.
This article is reviewed and published by the technical expert team of UA
Disclaimer: This article partially utilizes artificial intelligence assistance in its creation. If any content involves copyright or intellectual property issues, please let us know and we promise to verify and remove it as soon as possible.
Purchase recombinant protein, choose Nanjing UA-Bio
UA protein focuses on providing various protein reagents, raw materials, and services required for drug research and development, cell therapy, gene therapy, and basic scientific research, including drug target proteins, immune checkpoint proteins, cytokines, tool enzymes, customized protein expression, and full-length transmembrane protein development. Youai is committed to providing customers with high-quality products and professional services, and building a High-tech Biological Enterprise with International Competitiveness.
Target proteins | membrane proteins | cytokines | enzymes | viral antigens | protein customization
Nanjing UA Biotechnology Co., Ltd. Email:order@ua-bio.com Phone:+86-25-56221161
Product Information
Link:




©2021-2026 all rights reserved UA Biotechnology Co.,Ltd.
Privacy Policy 
网站备案号 浙ICP备2022019033号-2 浙公网安备33010202004867号
 Business license (three certificates in one)
Tel:+86-25-56221161
E-mail:order@ua-bio.com
All products sold on this website shall not be used for clinical diagnosis or treatment of humans or animals, and shall only be used for non-medical purposes such as industrial or scientific research.